Pharmacophore profiling highlights diverging binding modes of nemtabrutinib and ibrutinib across kinases
Kinase inhibition is a major pillar of targeted anticancer therapy, but rational control of kinase selectivity remains challenging because their ATP-binding pockets are highly homologous and conformationally dynamic. This can lead to off-target inhibition, toxicity, and reduced clinical effectiveness. Here, we present a computational workflow to rationalize selective compound activity by capturing subtle structural differences across kinases. We applied it to nemtabrutinib and ibrutinib, two BTK-targeting inhibitors with divergent activity in cell lines derived from activated B-cell-like (ABC) and germinal-center B-cell-like (GCB) diffuse large B-cell lymphoma (DLBCL). In vitro activity was correlated with in silico selectivity scores computed across 20 kinases relevant in lymphomas.
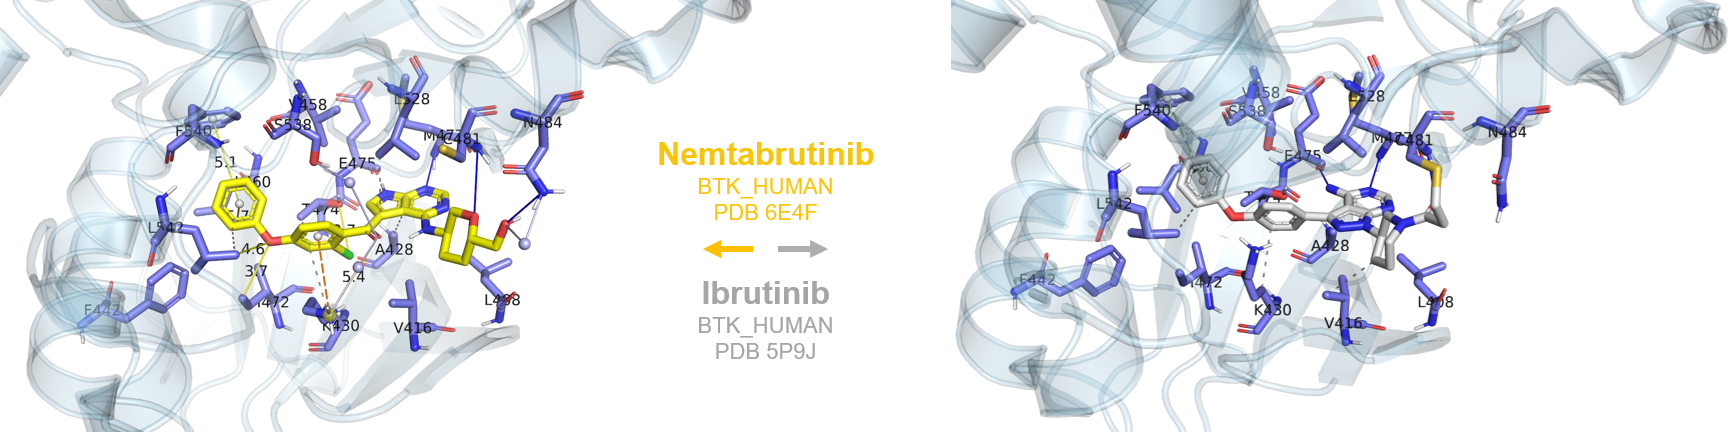
To enable scalable analysis of ATP-binding pocket variants, we collected metastable kinase conformations defined by DFG-motif geometry and associated dihedrals using AlphaFold2 templating and publicly available structures. Structural alignment of this conformational dictionary, followed by structure-based pharmacophore analysis, generated rich interaction maps of kinase pocket features. Compound fitting against these maps enabled pharmacophore matching and relative selectivity scoring, using BTK as the baseline confirmed target.
The analysis showed that the divergent profiles of nemtabrutinib and ibrutinib can be rationalized by the combined contribution of ten discriminant positions in the extended ATP-binding pocket, structurally aligned to BTK residues Q412, V416, F442, M449, L460, I472, T474, C481, N484, and L542. These positions modulate covalent reactivity, hydrogen bonding, hydrophobic packing, sulfur–aromatic interactions, π-stacking, cation–π contacts, and halogen-bonding opportunities. Interaction fingerprints highlighted FYN, FRK, MAST1, and SIK1–3 as candidate kinases preferentially compatible with nemtabrutinib over ibrutinib, providing a mechanistic hypothesis for differential anti-proliferative effects in GCB DLBCL models.
Overall, this proof of concept establishes a structure-guided strategy for interpreting selective compound activity across conserved kinase families. The workflow can also be extended toward automated contrastive screening and explicit design of selective kinase inhibitors optimized for desired target engagement while minimizing off-target interaction potential across the kinome.