Data-driven reactivity design through generative modeling with quantum chemistry validation
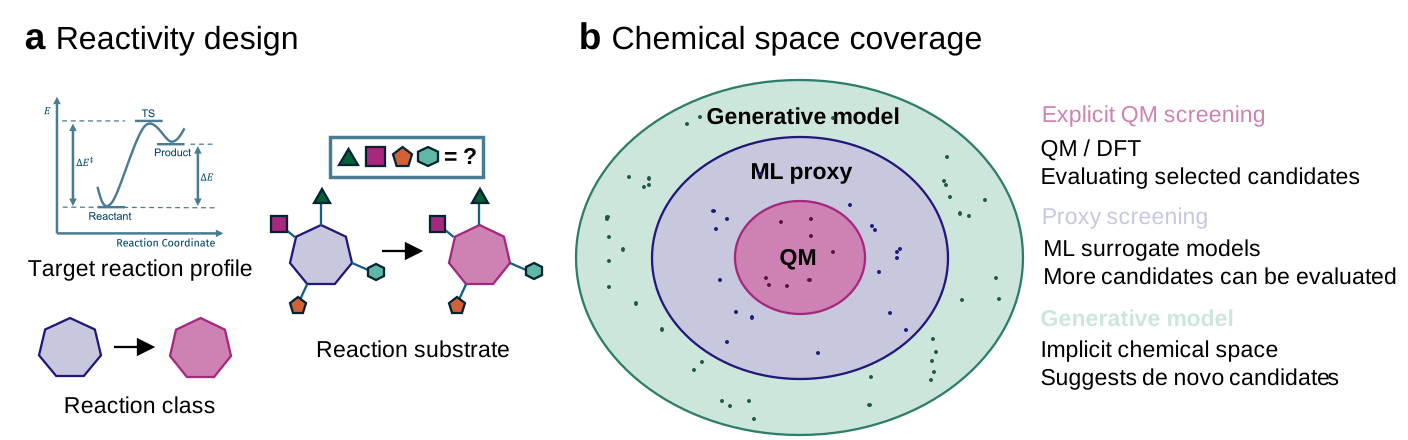
Data-driven inverse-design is a new emerging field in chemistry, showing promising results in drug discovery1,2,3 but remains underutilized in the context of reactivity design. This gap is largely attributed to data scarcity and the lack of generative models explicitly tailored for reaction-centric applications. While existing 3D generative frameworks possess the capacity for conditional generation of molecules, they cannot handle conditional generation on reactive properties, limiting their impact on reactivity design.4,5 In this work, we present a specialized generative modelling approach aimed at scaffold elaboration for reactivity design. Due to data scarcity, we limit ourselves in this study to reactivity design through scaffold decoration. We construct a curated dataset of around 10,000 DFT-level calculated keto-enol tautomerization reactions by subsampling a combinatorial library sharing a common N-Salicylideneaniline core scaffold, and additional 90’000 reactions calculated at xTB-level. We train a 3D structure-generating model on these datasets to produce transition state geometries that exhibit desired reactive properties. The inherent advantage of our approach over existing 2D approaches is easy validation of model predictions due to generated transition states. Our approach demonstrates the potential of targeted scaffold elaboration to advance computational reactivity design and provides a foundation for future developments in structure-based reaction modelling.
[1] Thomas E. Hadfield, Fergus Imrie, Andy Merritt, Kristian Birchall, Charlotte M. Deane, Journal of Chemical Information and Modeling, 2022, 62, 2280–2292.
[2] Xuhan Liu, Kai Ye, Herman W. T. van Vlijmen, Adriaan P. IJzerman, Gerard J. P. van Westen, Journal of Cheminformatics, 2023, 15, 24.
[3] Jaechang Lim, Sang-Yeon Hwang, Seokhyun Moon, Seungsu Kim, Woo Youn Kim, Chemical Science, 2020, 11, 1153–1164.
[4] Arne Schneuing, Charles Harris, Yuanqi Du, Kieran Didi, Arian Jamasb, Ilia Igashov, Weitao Du, Carla Gomes, Tom L. Blundell, Pietro Lio, Max Welling, Michael Bronstein, Bruno Correia, Nature Computational Science, 2024, 4, 899–909.
[5] Julian Cremer, Ross Irwin, Alessandro Tibo, Jon Paul Janet, Simon Olsson, Djork-Arné Clevert, arXiv, 2025, arXiv:2504.10564.