Molecular Dynamics Simulation of the Passivation Process of Hydroxylated Tin Oxide for Solar Cell Devices
Tin oxide (SnO2) semiconductor is frequently used in solar cell devices as an electron transport layer (ETL). However, due to its inherent highly reactive nature, interface passivation is required to ensure stability and prevent undesirable reactions with the active layer. In this context, the use of self-assembled monolayers (SAMs) with organic additives has been explored to protect the surface of the ETL.[1]
Notwithstanding, the molecular arrangement of these SAMs, as well as the reactive processes at the surface of SnO2, presents a high-dimensional problem that requires electronic structure methods with high accuracy and remains a challenge in the scientific community.[2]
In this work, we constructed a machine learning force field (MLFF) based on first-principle method to understand the reactive mechanisms involved in the hydroxylation process of SnO2 as well as the interactions between hydroxylated SnO2 and typical additives utilized in SAMs for surface passivation.
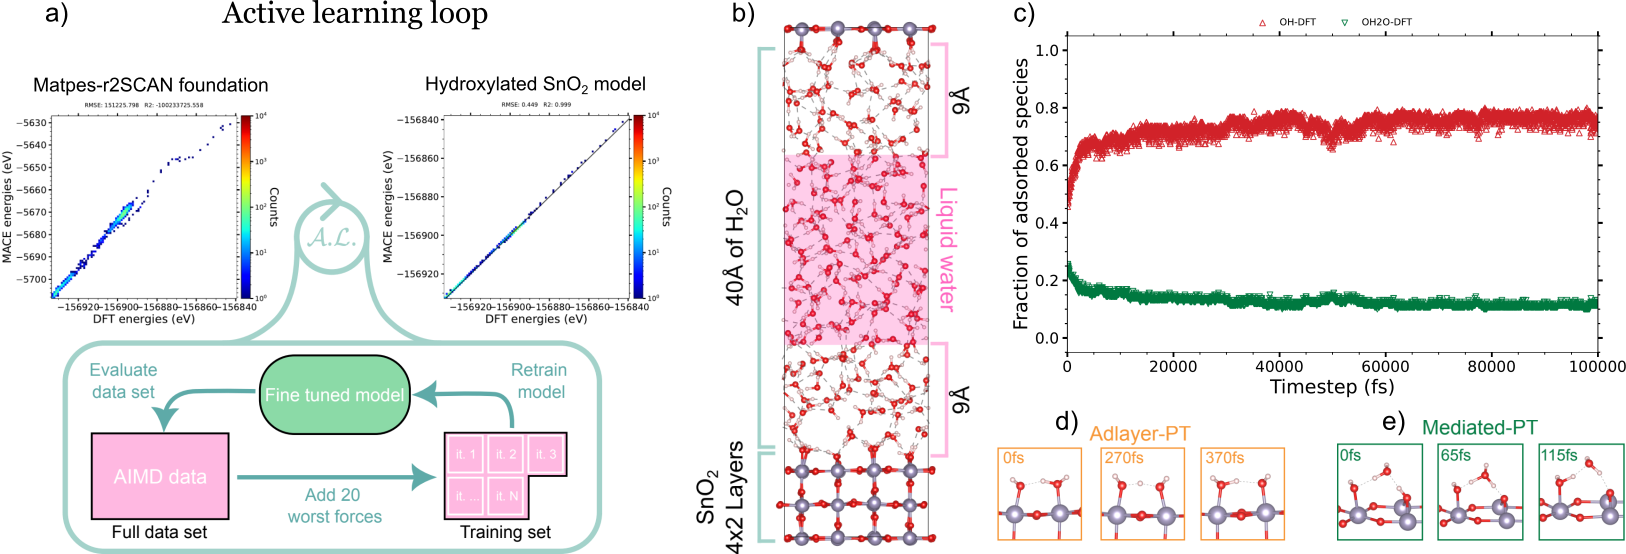
Figure 1: a) Representation of the active learning loop, b) representation of the simulated system, c) variation of hydroxylation of the surface (red) et adsorbed water (green) in function of simulation time, d) and e) mechanisms of surface proton exchange.
References:
[1] Park, S.M., Wei, M., Lempesis, N. et al. Nature, 624, (2023) 289–294
[2] M. Jia, Y.B. Zhuang, F. Wang, C. Zhang, J. Cheng, Precis. Chem., 2, 12, (2024) 644–654